Since the implementation of the Medical Device Regulation (MDR) in Europe, companies have had to maintain and execute a post-market surveillance (PMS) procedure. This systematic process involves collecting and analysing data on the performance and safety of medical devices after they have been released to the market. The primary goal is to help manufacturers identify and address potential issues that may arise during actual use, monitor the market for known risks, and identify new risks.
To achieve this, the PMS process interacts heavily with other processes, both in terms of data collection (input) and actions taken (output) – see figure 1.
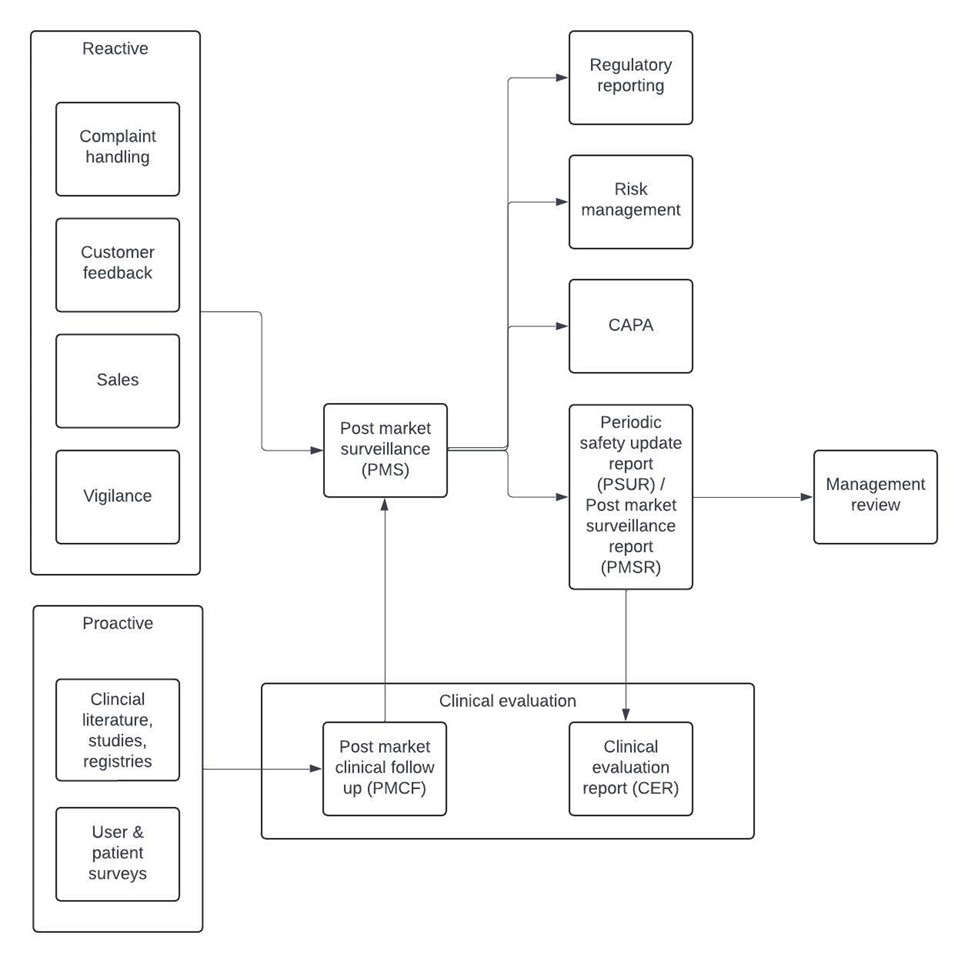
Figure 1: visualization of the information flow centered around the Post-Market Surveillance process.
The Need for a Common Language
Given the interdependence on a wide range of processes, a common language between these processes is essential. The easiest way to achieve this is by adopting the International Medical Device Regulators Forum (IMDRF) coding throughout all relevant processes in the company.
IMDRF coding (of which a partial example can be viewed in Table 1 below), is a coding system designed to streamline the reporting and analysis of medical device incidents. It supports effective and efficient monitoring of the performance and safety of medical devices. This standardization:
- Ensures that all stakeholders, including manufacturers, regulators, and healthcare providers, use consistent terminology, reducing misunderstandings and discrepancies in data interpretation and improving clarity in communication
- Minimizes the risk of errors and omissions in incident reports, leading to more precise and comprehensive data collection.
- Aligns with regulatory requirements: IMDRF terms are mandatory in the EU Manufacturer Incident Report (MIR) form for serious incident reporting under the MDR, and are also used by other regulators worldwide.
- Enables manufacturers to categorize and analyze incident data, identifying patterns and emerging issues faster and more efficiently.
- Ensures manufacturer PMS processes are aligned with global best practices.
Annex A (medical device problem) is only one of the IMDRF annexes. Annexes B to D describe the manufacturer’s cause investigation, Annexes E and F the clinical signs, symptoms and health impact for the patient and Annex G comprises the device component involved. Combining codes from several annexes gives a complete picture of an incident.
Table 1: Partial example of IMDRF Annex A. (source: Terminologies for Categorized Adverse Event Reporting (AER): terms, terminology and codes | International Medical Device Regulators Forum)
| Level 1 Term | Level 2 Term | Level 3 Term | Code | Definition |
| Manufacturing, Packaging or Shipping Problem | A02 | Problem associated with any deviations from the intended performance of the device that appear to relate to the manufacturing, packaging or shipping processes. | ||
| Product Quality Problem | A0201 | Problem associated with an inherent device characteristic that is not satisfactory as specified or delivered. | ||
| Dull, Blunt | A020101 | Problem associated with a device not being as sharp as intended or expected. | ||
| Nonstandard Device | A020102 | Problem associated with the device that does not meet the specifications or requirements for which it was manufactured (e.g. materials, parts, manufacturing process). | ||
| Packaging problem | A0205 | Problem associated with the materials used to construct the cover or outer wrapping of the device. | ||
| Device misassembled during manufacturing/shipping | A0206 | A device found incorrectly assembled when delivered to the user facility. | ||
| Device damaged prior to receipt by user | A0207 | Problem associated with packaging or shipping damage prior to receipt by the user. |
Extending IMDRF Coding Throughout the Company
Instead of using this coding solely for communication with external stakeholders, a much smoother process can be achieved by extending this “language” throughout the company. This would include:
- Logging complaints and feedback in categories aligned with IMDRF.
- Mapping the IMDRF coding to your risk management processes (e.g. hazards, hazardous situations and harms in ISO 14971), so that complaints can be traced back to the risk file.
- Reporting using the IMDRF coding in all subsequent processes (post-market surveillance report/Periodic safety update report, management review and clinical evaluation report).
Adopting a common language across the company would save time and improve clarity, ensuring that everyone is on the same page and working towards the same goals.
Simply writing the requirement into SOPs is, of course, not enough to ensure implementation. To implement such a change, we must plan training for the employees who will work with this IMDRF coding, and the integration into the existing data management systems. A common challenge is the resistance of employees who are used to working with the old system and don’t recognize the benefits the new system offers. The best way to get these employees on board would be to include them, from the start, in the implementation process, gather their feedback and provide continuous support to deliver a smooth transition.
Future trends
In conclusion, full IMDRF adoption not only delivers immediate efficiency gains but also prepares a PMS system for a future where data gathering will be increasingly AI-based. A common coding system throughout the various processes of the company allows for a smoother transition to a more automated way of working.